

The Long Read Sequencing Laboratory (LRS Lab) investigates the genetic basis of reproductive isolation and genome evolution and develops computational methods for analysing native DNA and RNA. Our research brings together Caenorhabditis genetics, open-source software development, bacterial and viral genomics, RNA biology and epigenomics. We combine genetic experiments, long-read sequencing and computational analysis to understand how genomes differ, how transcripts are organised and how DNA and RNA modifications contribute to biological variation. Laboratory Website | Software | GitHub
CityUHK Scholars | Current Openings
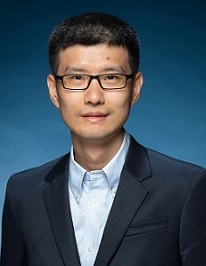
Prof. Runsheng Li is an Associate Professor in the Department of Infectious Diseases and Public Health at City University of Hong Kong. He received his veterinary degree from Northwest A&F University and his PhD from Nanjing Agricultural University. Before joining CityUHK, he worked at Hong Kong Baptist University.
His research focuses on the genetic basis of reproductive isolation, hybrid incompatibility and genome evolution, particularly using Caenorhabditis briggsae and Caenorhabditis nigoni as experimental models. His laboratory combines genetic crosses, introgression lines, experimental evolution and genomic analysis to investigate hybrid sterility, interactions between chromosomes and the evolutionary consequences of interspecific hybridisation.
He leads the Long Read Sequencing Laboratory, where computational method and open-source software development form a major, independent research programme. The group develops tools for nanopore signal analysis, DNA and RNA modification detection, transcript reconstruction, sequencing-quality assessment and pathogen-genome analysis. These methods support research across evolutionary genetics, bacterial and viral biology, and clinical methylomics.
We investigate how genetic differences between populations and species give rise to reproductive incompatibilities and shape genome evolution. Our principal experimental system is hybridisation between Caenorhabditis briggsae and Caenorhabditis nigoni. We combine controlled crosses, introgression lines, genetic mapping and functional analysis to identify the loci and molecular interactions responsible for hybrid male sterility. Particular interests include interactions between the X chromosome and autosomes, regulatory divergence and the disruption of gene expression in hybrids. We also use experimental hybrid evolution to investigate how parental genomic regions are retained or eliminated over generations. Our aims are to understand the contributions of selection, transmission-ratio distortion, recombination and transposable elements to introgression and the persistence of minority-parent ancestry. This research connects individual incompatibility loci with chromosome-scale genome evolution and the mechanisms that establish and maintain species boundaries. Algorithm and software development is a central research direction of the laboratory. We develop methods that turn long-read sequences and nanopore electrical signals into interpretable information about genomes, transcripts and nucleotide modifications. Our tools address modification-associated sequencing errors, comparative current-signal analysis, full-length transcript reconstruction, spliced-leader detection, long-read quality assessment and pathogen-genome analysis. Current development extends these capabilities towards scalable transcript and bacterial transcription-unit analysis, methylation-motif discovery and evidence-based methyltransferase–motif assignment. We aim to integrate sequence, protein structure, protein language models and genomic context while clearly separating computational predictions from experimentally established functions. We prioritise reproducible benchmarking, computational efficiency, transparent evidence and software that researchers can use and extend. We study bacterial and viral pathogens by integrating genome sequencing, transcriptomics, DNA methylation analysis and RNA modification profiling. For bacterial pathogens, our interests include methylation motifs, methyltransferases, restriction–modification systems and their variation across strains and environmental conditions. We investigate relationships between methylation, gene expression, mobile genetic elements and pathogen adaptation. For viral pathogens, we investigate genome variation, viral transcript organisation, genomic and subgenomic RNAs, RNA modification-associated signals and host responses to infection. Our work includes direct RNA sequencing of Sindbis virus and transmissible gastroenteritis virus, together with genome-analysis workflows and DNA methylation profiling for African swine fever virus. We use comparative controls, raw-signal evidence and complementary analyses to distinguish genuine biological variation from sequencing and analytical artefacts, without assuming that every predicted signal represents a confirmed chemical modification. We develop and apply long-read RNA sequencing methods to investigate transcript structure, RNA processing and epitranscriptomic variation. Our work encompasses full-length isoform discovery and quantification, alternative splicing, transcript boundaries, spliced-leader trans-splicing and operon organisation in nematodes and other eukaryotes, as well as bacterial and viral RNA populations. We compare native RNA with unmodified controls to identify and evaluate RNA modification-associated signals and connect transcript structure, abundance and modification within RNA populations. We develop and apply nanopore sequencing approaches to investigate DNA methylation in clinical research, particularly acute myeloid leukaemia and related haematological malignancies. Our work connects sequencing-method development with analysis of methylation patterns, disease-associated molecular variation and treatment-related changes. We aim to extend these approaches to low-input and circulating cell-free DNA, integrating methylation with sequence and fragment-level information for ageing and disease research. Priorities include sample preparation, sequencing quality, appropriate controls and the separation of biological variation from technical bias.Genetics of Reproductive Isolation and Hybrid Incompatibility
Computational Methods and Open-source Software Development
Pathogen Genomics and Epigenomics
Long-read Transcriptomics and RNA Biology
Clinical and Translational Methylomics
We develop open-source tools for genetics, long-read sequencing and pathogen analysis. Our software spans nanopore signal interpretation, transcript reconstruction, sequencing-quality assessment and reproducible genome-analysis workflows.
Hammerhead detects bacterial DNA modification-associated patterns using strand-specific nanopore sequencing mismatches. It supports de novo modification analysis and correction of modification-associated errors in bacterial genome assemblies. nanoCEM visualises and statistically compares nanopore current-event features at candidate DNA and RNA modification sites. It supports multiple preprocessing approaches and facilitates comparison between samples and controls. nanoSundial supports comparative nanopore signal analysis for RNA modification discovery. It was developed alongside our work on bacterial direct RNA sequencing and native-versus-unmodified RNA controls. trackcluster identifies and quantifies transcript isoforms from long-read RNA sequencing data. It was developed to investigate full-length transcript diversity during Caenorhabditis elegans development. trackcluster-rs provides a Rust implementation for scalable long-read transcript analysis. trackclusterTU extends our software development towards bacterial transcription-unit reconstruction and clustering of mapped long RNA reads. SLRanger detects spliced leaders and supports operon prediction from long RNA reads. It provides an integrated approach to analysing trans-splicing and transcript organisation in organisms such as nematodes. primerdiffer supports large-scale primer design for haplotype genotyping, connecting computational sequence analysis with experimental genetics. Giraffe processes and visualises long-read sequencing data, supporting assessment of read quality and systematic sequencing-error patterns across sequencing technologies. ANASFV is an integrated workflow for African swine fever virus whole-genome analysis, providing a reproducible framework for processing and characterising viral sequencing data.DNA and RNA Modification Analysis
Hammerhead
nanoCEM
nanoSundial
Transcriptomics and Genetics
trackcluster
trackcluster-rs and trackclusterTU
SLRanger
primerdiffer
Sequencing Quality and Pathogen Workflows
Giraffe
ANASFV
1. Tan, L., et al. (2026). Production of Abundant Defective Viral Genomes by an Insect-Specific Alphavirus Revealed by Nanopore Sequencing. Nature Communications. https://doi.org/10.1038/s41467-026-78770-9
2. Liu, X., et al. (2024). Nanopore strand-specific mismatch enables de novo detection of bacterial DNA modifications. Genome Research, 34(11), 2025–2038.
3. Tan, L., et al. (2024). Analysis of bacterial transcriptome and epitranscriptome using nanopore direct RNA sequencing. Nucleic Acids Research, 52(15), 8746–8762.
4. Lu, B., et al. (2025). Comprehensive comparison of the third-generation sequencing tools for bacterial 6mA profiling. Nature Communications, 16, 3982.
5. Shao, Y., et al. (2025). SLRanger: an integrated approach for spliced leader detection and operon prediction using long RNA reads. Briefings in Bioinformatics, 26(5), bbaf437.
Complete Publication Record — CityUHK Scholars
Laboratory Publications
The LRS Lab brings together experimental genetics, computational biology and long-read sequencing. We investigate biological questions while developing the methods needed to address them, connecting genetic experiments and sequencing data with algorithms, software and biological interpretation. Our research spans Caenorhabditis genetics and hybrid incompatibility, DNA and RNA modification analysis, bacterial and viral genomics, and long-read transcriptomics. Meet the Team | Explore the Laboratory We welcome applications from prospective PhD students interested in experimental genetics or computational method development. Applicants with prior research experience working with Caenorhabditis elegans will be given priority. Experimental projects focus on Caenorhabditis genetics and hybrid incompatibility. Computational projects focus on methods for nanopore DNA and RNA modification analysis. Applicants with backgrounds in genetics, molecular biology, bioinformatics, computational biology, computer science, statistics or related disciplines are encouraged to contact us. Programming experience is valuable for computational projects. Prospective applicants should send a CV and a brief description of their research interests. Computational applicants are encouraged to include a GitHub profile or examples of previous code. Current Openings and Application Information Prof. Runsheng Li Email: runsheng.li@cityu.edu.hk Telephone: +852 3442 7207 Office: Room 2-502, 5/F, Block 2, To Yuen Building, 31 To Yuen Street, Kowloon, Hong Kong Laboratory website: https://lrslab.github.io/ GitHub: https://github.com/lrslab/Long Read Sequencing Laboratory
PhD Opportunities
Contact
Department of Infectious Diseases and Public Health
City University of Hong Kong